Biomedical Engineer graduate of Universidad del Rosario, wet-lab enthusiast, and permanently curious person from Colombia 🇨🇴
I am happiest in the lab, where I often find myself hearing or asking:
“What happens if we try this?”
My path into biotechnology has moved through nanotechnology, mammalian cell culture, biomaterials, microbiology, and biological manufacturing.
One of the projects that made me fall in love with experimental biology involved testing nanoplatforms in mammalian cells, studying their cytotoxicity and trying to understand how engineered materials interact with living systems. Since then, my interests have kept expanding.
Lately, through my work, I have been spending much more time around microorganisms and I have become increasingly fascinated by how much these tiny living systems can tell us, produce and transform!
HTBAA feels like the perfect excuse to explore all of those worlds at once.
I am particularly interested in technologies that make advanced biotechnology more accessible to people and laboratories that do not have enormous amounts of infrastructure. I am also becoming increasingly curious about living systems as sensors: the idea that cells, microorganisms, or cell-free systems could not only manufacture something useful, but also detect, remember, or communicate information about their environment. That is one of the questions I want to explore through my HTBAA work.
Outside the lab
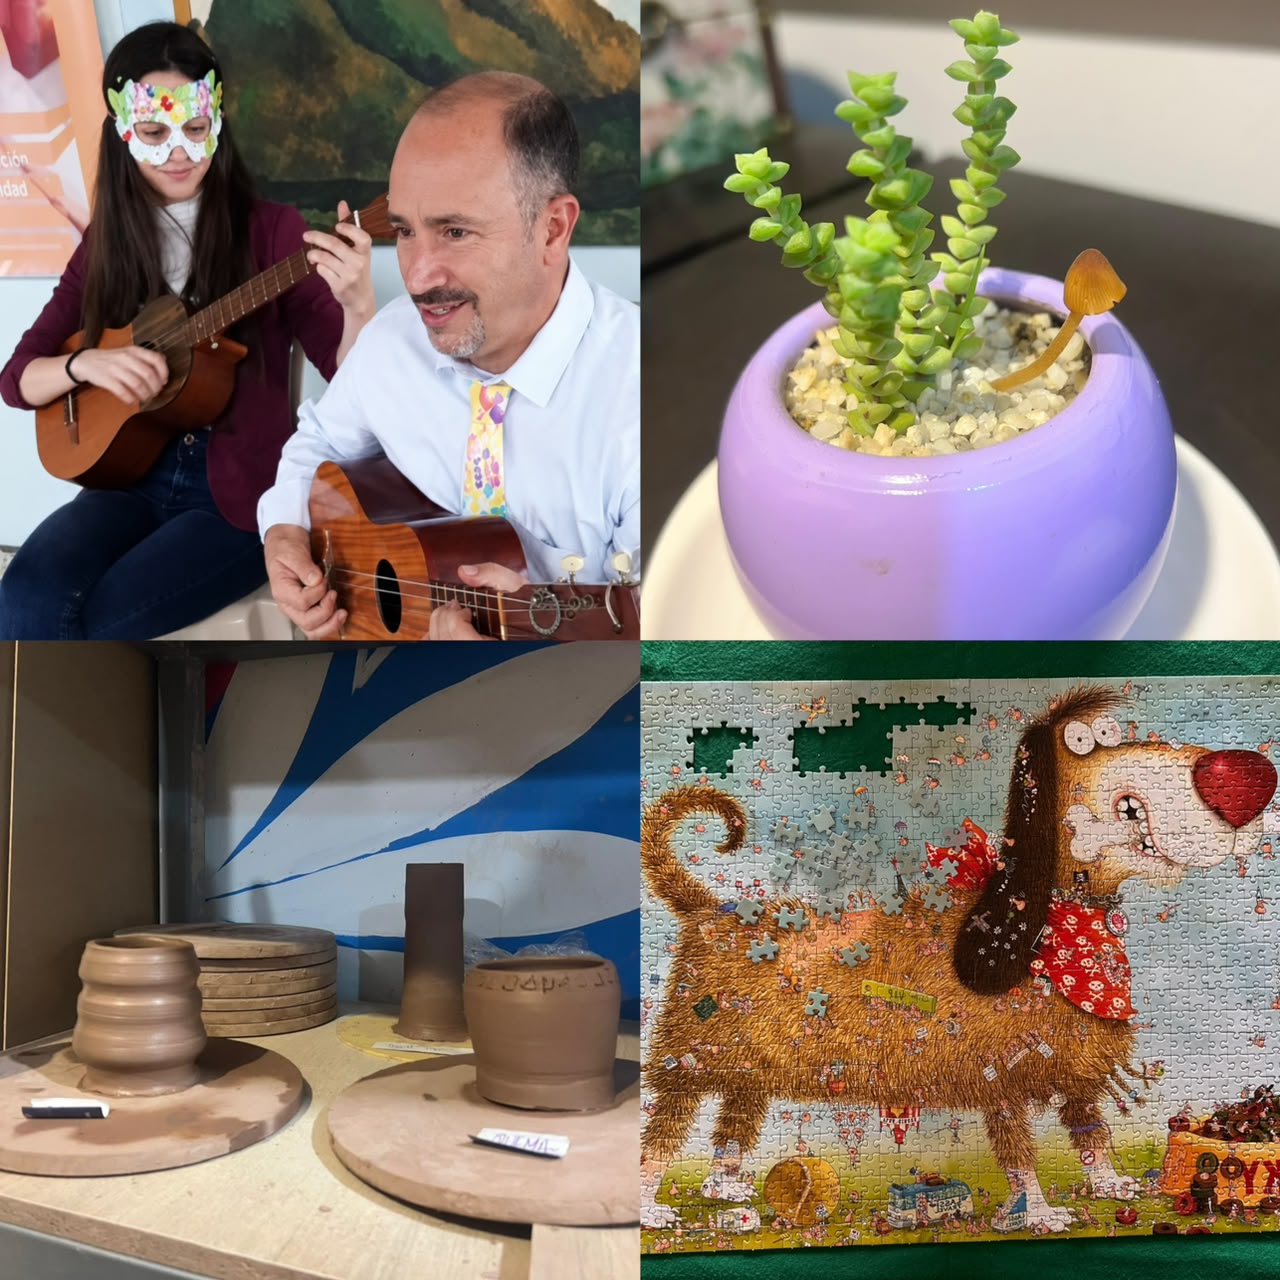
🎶 I play Colombian folk music
I play instruments with my dad, especially Colombian folk music such as llanera music and carranga. It is one of my favorite things we do together.
🍄 Biology also runs in the family
My dad is a biology teacher and an absolute fungi enthusiast. He can turn almost any walk into a conversation about the world of fungi.
I think some of his curiosity about living things definitely rubbed off on me.
🏺 I am learning ceramics
My first ceramic classes produced cups were… Let’s call them structurally experimental. You be the judge 🫣 I am getting better… Probably 😅.
🧩 I also love putting together huge jigsaw puzzles and watching anime
I really enjoy working on large jigsaw puzzles and watching anime in my free time.
What I hope to bring to this community
I want this website to show not only what I make, but how my way of thinking changes as I learn.
I really enjoy spaces where people come from completely different backgrounds but are willing to teach each other.
I hope to bring: curiosity, a strong willingness to learn, hands-on laboratory experience and, most importantly, the habit of trying to get the best out of both the experiments and the people around me.
I am here to learn things I do not know yet, discover new ways of thinking, and hopefully help other people do the same.
Say hi
Lina María Bello Rojas Biomedical Engineer · Bogotá, Colombia 🇨🇴
Subsections of <LINA MARIA BELLO ROJAS> — HTBAA Fall 2026
Week 2 HW
🧬 BioColdTrace — Homework #2
Part 1: Define your expression target
What exactly should the cells make?
In Homework #1, I proposed BioColdTrace, a biological Time–Temperature Indicator designed to transform cumulative thermal exposure into a persistent visible signal.
My original prototype was intentionally simple:
🦠 Yeast + nutrient source + colorimetric reporter
For this homework, I want to move from that general concept to a genetically defined output.
Instead of leaving the colorimetric response open, I would engineer Saccharomyces cerevisiae to produce a visible chromoprotein whose accumulation could eventually act as the readable output of the indicator.
1. 🎨 What molecule am I expressing?
The expression target I would use is amilCP, a blue-purple chromoprotein originally identified in the coral Acropora millepora.
Chromoproteins are particularly interesting for BioColdTrace because they produce a visible color without requiring fluorescence excitation or specialized instrumentation.
amilCP has an absorption maximum around 588 nm, giving it a strong blue-purple appearance. [1,2]
This makes it attractive for a system whose final output should ideally be readable simply by looking at it.
Why amilCP?
For BioColdTrace, I want the output to eventually satisfy several requirements:
👁️ Visible without specialized equipment
🧬 Genetically encoded
📈 Able to accumulate over time
📦 Compatible with a simple contained indicator
🔬 Still measurable quantitatively during development
amilCP fits these criteria particularly well because the reporter itself is colored.
This means the first engineered architecture could conceptually look like:
Importantly, amilCP would not itself be the temperature sensor.
It would be the output or recording molecule.
The mechanism determining when amilCP is expressed would be designed separately through the promoter and regulatory architecture.
This separation is useful because it allows me to independently optimize:
temperature sensing → gene regulation → visible output
The accumulation of a colored protein is also conceptually compatible with the idea of biological memory proposed in Homework #1.
If warmer conditions trigger reporter production, returning the system to refrigeration should slow further biological activity without immediately eliminating the protein that has already accumulated.
2. 🧬 What is the sequence?
The protein I would encode is amilCP from Acropora millepora.
For an actual DNA construct, I would use this protein sequence as the reference and generate a coding sequence codon-optimized for Saccharomyces cerevisiae.
Therefore:
Native coral sequence → protein reference Codon-optimized DNA sequence → yeast expression construct
The amino acid sequence would remain unchanged while the DNA sequence could be adapted to improve expression in the yeast chassis.
This distinction is important because the biological target is amilCP, while the exact nucleotide sequence used in the final construct can be engineered according to the codon preferences and design constraints of S. cerevisiae.
3. 🦠 What host or chassis?
I would keep Saccharomyces cerevisiae as the biological chassis proposed in Homework #1.
I originally selected yeast because:
it is inexpensive and easy to culture;
it grows on relatively simple media;
it is one of the best-characterized organisms in biotechnology;
extensive genetic and synthetic-biology tools are available;
its metabolism and growth are strongly affected by temperature;
it has already been explored in biosensing applications.
For the engineered version of BioColdTrace, I would narrow the chassis further and initially use:
Saccharomyces cerevisiae CEN.PK113-7D
CEN.PK113-7D is a haploid laboratory strain widely used in metabolic engineering, systems biology, and industrial biotechnology. Its genome has also been sequenced and characterized, making it a useful reference strain for quantitative engineering studies. [3]
Using a well-characterized strain is useful because I ultimately want BioColdTrace to establish a quantitative relationship between:
🌡️ temperature × ⏱️ exposure time × 🦠 cellular response × 🎨 reporter accumulation
A defined genetic background should make those relationships easier to study than starting with an undefined commercial yeast strain.
However, I would still consider the chassis an experimental variable.
If CEN.PK113-7D shows insufficient activity at refrigeration or intermediate temperatures, other S. cerevisiae strains adapted to lower temperatures could later be screened.
This would remain consistent with my original BioColdTrace proposal, where chassis selection itself was identified as one of the important questions to explore.
4. 📍 Where does the product need to go?
For the first BioColdTrace construct, amilCP would remain intracellular, primarily in the cytoplasm.
Therefore, I would not include a secretion signal peptide or organelle-targeting sequence.
The first architecture would simply be:
🦠 S. cerevisiae CEN.PK113-7D ↓ 🧬 intracellular expression of amilCP ↓ 🎨 accumulation of blue-purple color
Keeping the protein intracellular simplifies the first prototype because BioColdTrace does not require the reporter to perform any extracellular function.
Instead, the yeast cells themselves could act as small pigment-producing units.
There may also be an advantage for a future contained indicator: the reporter would remain associated with the biological material instead of needing to diffuse into the surrounding environment.
At this stage, I would therefore avoid adding unnecessary secretion or targeting sequences and keep the first construct as simple as possible.
🧠 What does this mean for BioColdTrace?
Defining amilCP solves one important part of the system:
What should the cells make?
The answer would now be:
A visible blue-purple chromoprotein whose accumulation can act as the readable output of BioColdTrace.
But this immediately creates the next — and more interesting — engineering question:
When should the cells make it?
The system should not simply become purple because yeast has been alive for a certain amount of time.
Instead, amilCP production should ultimately depend on thermal history.
For example, BioColdTrace should ideally be able to distinguish between conditions such as:
❄️ 48 h at 4 °C
and
🌡️ 4 h at 25 °C
even though both involve the passage of time.
The challenge is therefore not simply to express a colored protein.
The challenge is to create a regulatory architecture where temperature and exposure time determine how much reporter accumulates.
In that sense, the system can be separated into three modules:
🌡️ 1. Input
Temperature history
↓
🧬 2. Processing
Temperature-sensitive biological regulation
↓
🎨 3. Output
amilCP accumulation
This modular architecture is particularly useful because each component could potentially be optimized independently.
If amilCP works well as an output but the temperature-response mechanism does not, I could modify the regulatory system without replacing the reporter.
Likewise, if a better visible reporter becomes available later, the output module could be exchanged without completely redesigning the sensing mechanism.
I chose pTDH3 because it provides strong constitutive expression in S. cerevisiae, allowing the first construct to test whether amilCP can accumulate to a visually detectable level before introducing temperature-responsive regulation.
Important design decision
pTDH3 is not intended to be the final temperature-sensing element of BioColdTrace.
Unlike bacteria, yeast does not use a bacterial ribosome binding site or Shine–Dalgarno sequence.
Instead, translation initiation depends strongly on the sequence surrounding the ATG start codon.
A well-established Kozak-like sequence for S. cerevisiae is:
AAAAAAATGTCT
Because the ATG belongs to the coding sequence, I would define the separate 5′ UTR / Kozak part in Kernel as:
AAAAAA
and design the amilCP CDS to begin with:
ATGTCT...
When assembled, the junction becomes:
AAAAAAATGTCT...
||||||--- CDS ---
 5'UTR
Why this part?
I chose an A-rich yeast Kozak context to provide an efficient and host-appropriate translation initiation environment immediately upstream of the amilCP start codon.
3. 🎨 Coding sequence — amilCP
Part information
Part type in Kernel
CDS
Short name
amilCP_ySC
Protein
amilCP
Source organism
Acropora millepora
Reference nucleotide accession
AY646075.1
Reference protein accession
AAU06854
UniProtKB
Q66PV0
Final design
S. cerevisiae-codon-optimized CDS
The coding sequence would encode the same amilCP protein defined in Part 1.
The natural amilCP sequence originates from the coral Acropora millepora and is available as GenBank AY646075.1 / AAU06854.
For my final Kernel part, however, I would create a derived DNA sequence codon-optimized for S. cerevisiae while preserving this amino acid sequence.
Therefore:
AAU06854 protein sequence ↓ codon optimization for S. cerevisiae ↓ amilCP_ySC CDS
The CDS must include:
an ATG start codon;
the complete amilCP open reading frame;
a stop codon.
Sequence note
The exact nucleotide sequence recorded here should be the final codon-optimized DNA sequence generated/imported in Kernel, rather than a manually reverse-translated sequence.
Final Kernel sequence:
[PASTE FINAL amilCP_ySC CODON-OPTIMIZED SEQUENCE EXPORTED FROM KERNEL HERE]
This ensures that the sequence documented on the website is identical to the sequence actually used in the genetic design.
Why this part?
I chose amilCP because it produces a directly visible blue-purple chromoprotein, allowing reporter accumulation to be observed without fluorescence excitation or specialized instrumentation.
4. 📍 Targeting — none
Part information
Part type
Signal peptide / targeting sequence
Part used
None
Destination
Cytoplasm
No secretion signal, transmembrane domain, or organelle-targeting peptide would be added.
amilCP does not need to leave the cell to perform its role in BioColdTrace.
Instead:
yeast cell → intracellular amilCP accumulation → visible cellular color
Keeping the protein cytoplasmic also makes the initial construct simpler and avoids introducing secretion efficiency as another experimental variable.
Why no targeting sequence?
No targeting sequence is required because amilCP is intended to accumulate intracellularly as the visible reporter, and secretion would add unnecessary complexity to the first prototype.
5. 🛑 Transcription stop — CYC1 terminator
Part information
Part type in Kernel
Terminator / 3′ UTR
Short name
tCYC1
Organism
Saccharomyces cerevisiae
Source
CYC1 3′ region
Function
Transcription termination and mRNA 3′-end formation
I would use the S. cerevisiae CYC1 terminator.
CYC1-derived 3′ sequences are widely used in yeast expression vectors and provide the signals required for proper transcript termination and 3′-end processing.
I chose the CYC1 terminator because it is a well-established S. cerevisiae 3′-end element that provides reliable termination and transcript processing in yeast expression constructs.
6. ⭕ Vector context — pRSII41K
Instead of designing a yeast plasmid backbone from scratch, I would reuse an existing backbone.
Selected backbone
pRSII41K / pRSII41K-TDH3pr
Addgene plasmid #194522 / #194528
The particularly convenient version for this project is:
pRSII41K-TDH3pr — Addgene #194528
because it already contains:
TDH3 promoter → MCS → CYC1 3′ UTR
Part information
Part type in Kernel
Backbone
Short name
pRSII41K
Yeast replication
CEN6 / ARS4
Yeast copy number
Low-copy
Yeast selection
kanMX → G418 resistance
E. coli replication
pBR322 origin
E. coli selection
bla → ampicillin resistance
Source
Addgene #194522 / #194528
The backbone uses the CEN6/ARS4 system, producing a relatively low-copy plasmid in yeast.
For BioColdTrace, I prefer a low-copy backbone over a high-copy 2µ plasmid for the first quantitative experiments because I ultimately want to compare:
temperature → biological activity → reporter accumulation
and reducing plasmid copy-number effects should make expression easier to interpret.
The kanMX marker also provides G418 resistance for selection of transformed yeast.
Why this backbone?
I chose pRSII41K because it is a low-copy S. cerevisiae expression backbone with convenient G418 selection, and an existing version already provides the TDH3 promoter and CYC1 3′ UTR needed for this expression cassette.
🧩 Final part list
Layer
Kernel part type
Part
Main function
Transcription start
Promoter
pTDH3
Strong constitutive transcription
Translation start
5′ UTR
yKozak_A6
Efficient yeast translation initiation
Coding sequence
CDS
amilCP_ySC
Blue-purple visible output
Targeting
Signal peptide / tag
None
Protein remains cytoplasmic
Transcription stop
Terminator
tCYC1
Transcript termination and 3′ processing
Vector context
Backbone
pRSII41K
Low-copy yeast replication + G418 selection
🧬 Final construct
The first BioColdTrace genetic construct can therefore be summarized as:
🌡️ Why is the promoter constitutive if BioColdTrace should sense temperature?
This is an intentional design decision.
The final BioColdTrace system should ultimately make amilCP accumulation dependent on thermal history, but I first need to characterize the reporter independently of that sensing mechanism.
The first construct therefore asks:
Can my selected yeast chassis produce enough amilCP to generate a measurable and visible output?
If the answer is yes, this construct provides a baseline response.
A later version can replace or regulate the constitutive transcriptional input with a temperature-responsive mechanism:
BioColdTrace v1

constitutive expression
        ↓
Does the reporter work?
        ↓
amilCP characterization
        ↓
baseline
This modular approach lets me debug the system one layer at a time rather than trying to determine simultaneously whether a failure comes from the temperature sensor, the regulatory architecture, or the chromoprotein reporter.
Ultimately, the goal remains:
thermal history → biological computation → accumulated visible memory
💻 What I would create in Kernel
I would create or import the following records:
pTDH3
Type: Promoter
Source: S. cerevisiae TDH3 promoter
yKozak_A6
Type: 5′ UTR
Sequence: AAAAAA
amilCP_ySC
Type: CDS
Source protein: AAU06854 / Q66PV0
Designed: codon-optimized for S. cerevisiae
tCYC1
Type: Terminator
Source: S. cerevisiae CYC1 terminator
pRSII41K
Type: Backbone
Source: Addgene #194522 / #194528
I would then assemble them in the following order:
pTDH3 → yKozak_A6 → amilCP_ySC → tCYC1
inside the pRSII41K backbone.
📚 References
[1] Alieva, N. O., Konzen, K. A., Field, S. F., et al. (2008). Diversity and evolution of coral fluorescent proteins. PLoS ONE, 3(7), e2680. https://doi.org/10.1371/journal.pone.0002680
[5] Dao Duc, K., Song, Y. S. (2018). The impact of ribosomal interference, codon usage, and exit tunnel interactions on translation elongation rate variation. PLoS Genetics. [Used here together with published analyses of the S. cerevisiae Kozak-like initiation context.]
[6] Osborne, B. I., & Guarente, L. (1989). Mutational analysis of a yeast transcriptional terminator. Proceedings of the National Academy of Sciences, 86(11), 4097–4101. https://doi.org/10.1073/pnas.86.11.4097
[7] Smith, J. R., et al. (2024). Auxotrophy-Independent Plasmid Shuttle Vectors for Applications in Diverse Yeasts. Applied Microbiology, 4(1), 453–469. https://doi.org/10.3390/applmicrobiol4010031
🧬 Week 1 Homework — Two Ideas, One Goal: Biomanufacturing for Real-World Impact
Exploring how biology can help solve practical challenges in biotechnology and healthcare.
For my first HTBAA assignment, I found myself thinking about two different ideas that connect my background in Biomedical Engineering, my professional experience, and my interest in making biotechnology more accessible.
Rather than choosing just one at this early stage, I wanted to share both. Although they address very different challenges, they are driven by the same question:
How can we use living systems to develop affordable, practical, and meaningful solutions for problems that matter?
🌱 My Two Project Ideas
🌡️ Idea 1 — BioColdTrace
A biological time–temperature indicator for cold-chain monitoring
Could a tiny biological system reveal whether a temperature-sensitive product has experienced harmful thermal exposure?
Inspired by my experience working with fetal bovine serum, this project explores using microorganisms to transform cumulative temperature exposure into a persistent, visible signal.
Bacteriophage-based wound dressings for rural communities in Colombia
Could bacteriophages be incorporated into affordable wound dressings to help control bacterial infections and explore alternatives to conventional antimicrobial treatments?
Inspired by the challenges of accessing healthcare in rural Colombia and my node’s experience working with bacteriophages, this project explores combining phage biomanufacturing with biocompatible materials for localized wound care.
BioColdTrace connects directly with my professional experience in biotechnology and quality control, while PhageGuard reflects my interest in biomedical innovation, microbiology, and developing healthcare solutions with a potential social impact in Colombia.
Both ideas are still exploratory, and I see HTBAA as an opportunity to understand their scientific feasibility, investigate possible biological manufacturing systems, and learn how automation could help transform an initial concept into a reproducible biomanufacturing process.
Two different ideas, but one shared motivation: making biotechnology useful, accessible, and relevant to real-world needs.
🌡️ Project 1 — BioColdTrace
Can a tiny living system reveal what happened before a product reached us?
1. 🌍 What would I like to biomanufacture, and why?
What if a product could carry a small biological memory of what happened to it before it reached us?
I would like to develop BioColdTrace, a low-cost biological Time Temperature Indicator (TTI) designed to convert cumulative thermal exposure into a persistent and visible signal.
The idea is partly inspired by my work with fetal bovine serum (FBS), a widely used component in biotechnology for which temperature control during transportation, processing, and storage is essential. A material can arrive cold while still having experienced an important temperature excursion earlier in its journey.
BioColdTrace would not try to reconstruct the entire temperature history. Instead, it would try to answer a simpler question:
Has enough thermal exposure accumulated to trigger a meaningful warning?
The concept could eventually be adapted to many cold-chain products after product-specific calibration:
🧪 Biotechnology: Enzymes, antibodies, media components, and other sensitive reagents. 🧬 Diagnostics: Samples moving from collection points to centralized laboratories. 🥛 Food: Dairy, meat, seafood and other refrigerated products. 🐄 Veterinary and rural supply chains: Temperature-sensitive biological products transported over long distances. 💉 Biologics and vaccines: As a complementary indicator where validated for a specific product. 🎓 Education & community biotech: An accessible system for teaching metabolism, sensing and biomanufacturing.
Existing time–temperature indicators have already been applied across food and cold-chain monitoring contexts [4], while Vaccine Vial Monitors provide a well-established example of cumulative heat-exposure indication for vaccines [5].
This makes the question behind the project especially interesting to me:
Who gets access to quality information when sophisticated monitoring cannot be placed on every individual product?
How would it compare with what already exists?
📊 Electronic data logger
🏷️ Existing TTIs
🦠 BioColdTrace — proposed
Exact temperature history
✅ Yes
❌ Usually no
❌ No
Cumulative thermal exposure
Calculated from data
✅ Yes
✅ Intended
Visual result without software
Usually no
✅ Often yes
✅ Intended
Battery/electronics required
Usually yes
Usually no
No
Can travel with individual products
Possible, but adds cost/logistics
✅ Yes for some TTIs
✅ Design goal
Product-specific calibration
Digital thresholds can be configured
✅ Already possible
✅ Biological tuning would be explored
Cost profile
Higher upfront cost; may be reusable depending on the device
Varies widely depending on technology and supplier
Designed for low-cost production using simple biological components; cost still to be validated
Main advantage
Complete quantitative history
Simple cumulative indication
Potentially open, biologically tunable, and locally manufacturable
Importantly, existing TTIs already solve several of these problems very well. So BioColdTrace only becomes interesting if biology adds something genuinely useful.
My hypothesis is that its value could come from four characteristics:
🎛️ Tunable — changing strain, cell concentration, nutrients, or reporter chemistry could modify response kinetics.
🔓 Open — the formulation could potentially be reproduced and modified using accessible biotechnology infrastructure.
📦 Unit-level — a small, inexpensive indicator could travel with individual packages.
🧬 Biological — rather than simply recording temperature, the system would use biological kinetics to integrate time and temperature into one response.
The goal is therefore not to make a cheaper thermometer. It is to explore whether biology can provide a useful new way of making cold-chain information accessible.
2. 🦠 What living system would I use?
My starting chassis would be the yeast Saccharomyces cerevisiae.
I chose it for practical and biological reasons:
it is inexpensive and easy to culture;
it grows on relatively simple media;
it is one of the most extensively characterized microorganisms in biotechnology;
many genetic and synthetic-biology tools already exist for it;
its growth and fermentation kinetics are strongly affected by temperature;
yeast has already been explored as a chassis for biological sensing. [3–5]
Published work shows that S. cerevisiae grows and ferments substantially more slowly at low temperatures than around its preferred growth range. That difference is exactly what I would try to exploit. [4]
How could BioColdTrace work?
The first prototype could be deliberately simple:
🦠Yeast + nutrient source + colorimetric reporter
→ ❄️ Cold: slow metabolism → 🌡️ Warm excursion: metabolism accelerates → 🧪 Biochemical change accumulates → 🎨 Color signal appears and remains
The key feature I would look for is memory.If a product is exposed to warmer conditions and is later returned to refrigeration, the signal should not simply disappear. Cooling should slow further activity, while the change already generated should remain visible.
Possible readouts could include changes in pH, redox state, or another metabolically generated signal coupled to a colorimetric reporter. I would not assume in advance which mechanism is best; comparing those possibilities would be part of the project.
I also do not want to assume that S. cerevisiae is necessarily the final chassis.
If its response is too slow, too variable, or poorly matched to relevant cold-chain conditions, another microorganism or even a cell-free system could ultimately be more appropriate.
That makes chassis selection itself one of the most interesting parts of the project.
And, on a more personal note, starting with a fungus feels fitting because my dad is a biology teacher and a lifelong fungi enthusiast I know I can count on his curiosity, enthusiasm, and support as I explore this project.
4. 🤖 How would I use automation?
For BioColdTrace, I see automation mainly as a way to move from one formulation that seems to work to a systematically tested biological system.
I would combine three different environments:
🧪 Local experimentation
At my workplace, I already have access to basic laboratory infrastructure including microbial culture, incubation, micropipettes, centrifugation, pH measurement, absorbance/transmittance measurements, and standard laboratory equipment.
This gives me a realistic way to begin testing questions such as:
Does yeast produce a measurable temperature-dependent response?
Which reporter gives the clearest signal?
Does the signal remain after the system returns to refrigeration?
How reproducible is the response?
In other words:
My local laboratory can help me ask: “Does it work?”
🤖 HTBAA remote automation
As a Committed Listener, I would also have the opportunity to design experiments that can be executed remotely through the automation infrastructure used in the HTBAA labs.
The course specifically describes global students designing experiments, receiving assigned wells in automated plates, having experiments executed at MIT or Ginkgo, and receiving the resulting data back. Multiple experimental rounds may also be possible.
I would use that experience to learn how to translate biological questions into robotic protocols and systematic screening workflows.
For BioColdTrace, a screening matrix could compare variables such as:
Rather than changing one variable at a time, automation could make it possible to compare many combinations with greater reproducibility. Where compatible instruments are available, automated optical measurements could also help generate quantitative response curves.
💻 From experiments to decisions
The resulting data could feed into a simple Python/Jupyter workflow:
This would allow each experimental round to inform the next one.
The course also introduces desktop liquid-handling robots, Cloud Labs, and programmable microfluidics, so later in the semester I would like to explore whether miniaturization could reduce reagent volumes or eventually allow several biological response formulations to be incorporated into one indicator.
🌍 Where the node becomes important
What I would value most is having a community and instructors who can help me decide:
Which biological variables are worth testing? Which chassis or reporter makes the most sense? What should I automate first? What result would actually be scientifically meaningful?
Depending on the node’s capabilities, additional local experimentation may also be possible, but I know that course does not guarantee that for every project.
So I see the three roles as complementary:
Local lab → Does it work? HTBAA automation → How can I test many versions systematically? Node → Which questions and design choices are most worth pursuing?
5. ❓ What don’t I know yet?
At this stage, the most important unknowns are not small technical details. They are the questions that will determine whether BioColdTrace is actually worth pursuing.
1. Can a biological response be matched to a meaningful cold-chain failure?
A color change alone is not enough. The response kinetics would eventually need to correspond to the thermal sensitivity of a real product.
For example:
Should the indicator respond to several hours at room temperature? A short high-temperature excursion? Repeated smaller excursions?
I do not yet know how closely a biological response can be tuned to those different thermal histories. This is probably the most important calibration problem in the project.
2. Is S. cerevisiae actually the right biological system?
Yeast is an attractive starting chassis because it is accessible, inexpensive, and well characterized.
But I still need to determine whether it can provide:
sufficient activity at relevant temperatures;
a clear difference between cold and warm conditions;
reproducible kinetics;
a persistent and easily readable signal.
If it cannot, another microorganism, an engineered strain, or a cell-free system may ultimately be more appropriate.
The microbial manufacturing, cell-free, genetic circuit, and non-model organism modules of HTBAA could help me evaluate those alternatives throughout the semester.
3. Can the system become a practical indicator rather than only an interesting biological experiment?
Even if the biology works, a real indicator would still need:
I am particularly interested in learning whether tools introduced later in the course — including automated screening, measurement, and microfluidics — could help address some of these challenges.
And behind all three questions is an even larger one:
Does biology provide enough additional value to justify using it instead of an existing chemical TTI or electronic data logger?
BioColdTrace would only be worth developing if its biological nature creates a meaningful advantage, whether through tunability, accessibility, open manufacturing, or the ability to match specific thermal-response profiles.
I want to know whether biology can turn thermal history into useful, accessible, and actionable information.
That is the question I hope to explore throughout HTBAA.
📚 References
[1] Liu, Y., Li, L., Yu, Z., Ye, C., Pan, L., & Song, Y. (2023). Principle, development and application of time–temperature indicators for packaging. Packaging Technology and Science, 36(1 0), 833–853. https://doi.org/10.1002/pts.2761
[3] Kavšček, M., Stražar, M., Curk, T., Natter, K., & Petrovič, U. (2015). Yeast as a cell factory: current state and perspectives. Microbial Cell Factories, 14, 94. https://doi.org/10.1186/s12934-015-0281-x
[4] García-Ríos, E., Morard, M., Parts, L., Liti, G., & Guillamón, J. M. (2017). The genetic architecture of low-temperature adaptation in the wine yeast Saccharomyces cerevisiae. BMC Genomics, 18, 159. https://doi.org/10.1186/s12864-017-3572-2
Could we use bacteriophages to make wound care more accessible in rural Colombia?
1. 🌍 What would I like to biomanufacture, and why?
What if a simple wound dressing could do more than cover an injury? What if it could use biology to help fight harmful bacteria?
I would like to develop PhageGuard, a potentially affordable wound dressing incorporating bacteriophages into a biocompatible material, such as a hydrogel, to explore a new approach to localized bacterial infection control.
The idea is inspired by two things that I find particularly meaningful: my interest in biomedical engineering and biotechnology, and the healthcare challenges faced by rural communities in Colombia.
In agricultural environments, farmers and rural workers may experience cuts, abrasions, and other injuries during their daily activities. Exposure to soil, water, animals, and agricultural equipment can introduce microorganisms into wounds.
Although many minor injuries heal without complications, some can become infected. When access to healthcare is limited by distance, transportation, or economic barriers, obtaining timely assessment and treatment may be difficult.
This made me think about a question:
Could we use biotechnology to develop more accessible wound-care materials for communities where healthcare resources are limited?
🦠 Why bacteriophages?
Bacteriophages, or simply phages, are viruses that infect bacteria.
Unlike broad-spectrum antibiotics, individual bacteriophages generally recognize specific bacterial hosts. Some phages follow a lytic cycle, infecting susceptible bacteria and ultimately causing their destruction.
This specificity is scientifically interesting because it creates the possibility of targeting particular bacterial pathogens while potentially limiting effects on unrelated microorganisms.
However, it also introduces an important limitation: a bacteriophage that is effective against one bacterial strain may not work against another.
Bacteriophages are being investigated as potential tools against antibiotic-resistant bacterial infections. The World Health Organization identifies antimicrobial resistance as a major global public health challenge, including resistance in pathogens such as Pseudomonas aeruginosa and Staphylococcus aureus [1].
My idea would be to explore whether selected lytic bacteriophages could be incorporated into a wound dressing while maintaining their biological activity.
Importantly, I would not assume that PhageGuard could prevent every wound infection, replace antibiotics, or safely treat wounds without medical evaluation.
Instead, I would investigate whether a phage-loaded material could eventually become a targeted, complementary antimicrobial technology, following appropriate preclinical and clinical validation.
🩹 What exactly would PhageGuard be?
The proposed product would combine three components:
1. 🧬 Bacteriophages
Selected lytic phages capable of recognizing bacterial strains relevant to wound infections.
2. 🧪 A biocompatible material
A hydrogel or another suitable wound-dressing matrix that could protect the phages, support their stability, and potentially enable localized release.
3. 📦 An accessible dressing format
A practical, individually packaged material designed with affordability, portability, storage stability, and ease of use in mind.
The long-term goal would be a dressing that could complement existing wound-care strategies in settings where access to advanced healthcare may be limited.
How would it compare with existing approaches?
Feature
🩹 Conventional dressing
🧪 Antimicrobial dressing
🦠 PhageGuard — proposed
Physical wound protection
✅ Yes
✅ Yes
✅ Design goal
Targeted action against specific bacteria
❌ Usually no
Varies by technology
✅ Intended
Uses biological antimicrobial agents
❌ No
Depends on formulation
✅ Yes
Active against antibiotic-resistant strains
Not inherently
Depends on agent and organism
Potentially, if the phage is active against the strain
Broad antimicrobial coverage
❌ No
Possible
Limited by phage host range
Requires identification of target bacteria
No
Usually not
May be necessary
Low-cost manufacturing
Possible
Depends on technology
Design goal; not yet demonstrated
Clinical validation
Established for approved products
Product-specific
Still required
Importantly, antimicrobial wound dressings already exist, including products based on silver, iodine, and other antimicrobial technologies.
So PhageGuard would not be interesting simply because it is antimicrobial.
Its potential value would come from combining biological specificity, biomaterials, and potentially adaptable manufacturing.
Why is this idea particularly interesting to me?
Beyond its potential application in rural Colombia, PhageGuard connects several areas that I want to explore more deeply:
🧬 Microbiology and bacteriophages: Understanding how viruses interact with bacterial hosts.
🩹 Biomedical engineering: Designing functional biomaterials for healthcare applications.
🤖 Automation: Using systematic experimentation to compare materials, formulations, and biological performance.
🌱 Accessible biotechnology: Exploring whether advanced biological technologies can eventually be adapted to communities with limited healthcare resources.
🤝 My HTBAA node: My node has experience working with bacteriophages, which makes this project especially exciting. I would love to learn from that experience and explore whether it could help transform the idea into a scientifically meaningful research project.
Ultimately, the question that motivates me is:
Can we make sophisticated biological technologies useful in places where sophisticated healthcare infrastructure is not always available?
2. 🦠 What living system would I use?
My proposed biological manufacturing system would be a bacteriophage–bacterial host platform.
Unlike BioColdTrace, where a microorganism directly generates a measurable response, PhageGuard would use microorganisms to manufacture the active biological component of a biomedical material.
🧬 The biological chassis
The central manufacturing system would involve a suitable bacterial host and a compatible bacteriophage.
The bacterium would act as the manufacturing chassis.
The bacteriophage would be the desired biological output.
This distinction is important because bacteriophages cannot reproduce independently. They depend on susceptible bacterial cells to produce new viral particles.
At a conceptual level, the process would involve:
🦠 Suitable bacterial host
↓
🧬 Compatible lytic bacteriophage
↓
🔬 Phage replication inside bacterial cells
↓
🧪 Recovery and purification of phage particles
↓
🩹 Incorporation into a biocompatible dressing
↓
📦 PhageGuard prototype
The manufacturing process would need to include appropriate quality control, removal of bacterial impurities, and verification of phage identity and activity.
Which bacteriophages would I consider?
I would initially investigate the scientific and clinical relevance of phages targeting bacteria commonly associated with wound infections.
Two potential research targets are:
🧫 Staphylococcus aureus
A clinically important Gram-positive bacterium associated with skin and soft-tissue infections. Certain strains, including methicillin-resistant S. aureus (MRSA), present important treatment challenges.
🧫 Pseudomonas aeruginosa
A Gram-negative opportunistic pathogen associated with difficult-to-treat wound infections, including burns and chronic wounds. Some strains demonstrate resistance to multiple antibiotics.
Both organisms appear in the WHO’s 2024 priority pathogens framework [1].
However, I would not attempt to develop a broad phage cocktail against multiple pathogens immediately.
My first research objective would be much narrower:
Can a suitable bacteriophage remain biologically active after being incorporated into a wound-dressing material?
A single well-characterized, non-engineered lytic phage could be sufficient for investigating that question in an appropriately equipped and authorized research environment.
The final choice of phage and host would depend on strain characterization, biosafety requirements, available expertise, and the capabilities of the collaborating node.
🧪 Why use a hydrogel?
Hydrogels are three-dimensional polymer networks capable of retaining substantial amounts of water.
They are particularly interesting in biomedical engineering because their properties can be adjusted for different applications, including wound dressings and controlled delivery systems.
For PhageGuard, a hydrogel could potentially serve several functions:
Provide a physical matrix for incorporating bacteriophages.
Help maintain an environment compatible with phage stability.
Allow investigation of how phages are retained or released.
Support the development of a flexible material appropriate for wound-care applications.
Potential material families worth investigating include alginate-based and other biocompatible hydrogel systems.
However, not every hydrogel is compatible with every bacteriophage.
Polymer chemistry, pH, charge, processing conditions, and storage environment may affect phage activity.
Published research has already explored phage-containing hydrogels and other topical delivery systems, with promising preclinical findings but important remaining challenges in formulation, stability, and clinical translation [2,3].
How could PhageGuard work?
The proposed mechanism would be:
🩹 Phage-loaded wound dressing
→ 💧 Contact with an appropriate local environment
→ 🧬 Release or availability of viable phages
→ 🦠 Encounter with susceptible target bacteria
→ 🔬 Phage-mediated bacterial infection and lysis
→ 📉 Potential reduction in susceptible bacterial populations
This is a proposed mechanism, not a demonstrated therapeutic outcome for PhageGuard.
An important distinction is that phages would not manufacture the hydrogel itself.
Instead, the project would combine two manufacturing elements:
Biomanufacturing: Producing bacteriophages using biological systems.
Biomaterial fabrication: Incorporating the purified biological product into a suitable dressing matrix.
I find this particularly interesting because it brings microbial manufacturing and biomedical materials together in a single product concept.
3. 📦 How much would I need to make, and how long would it take?
For this assignment, I would define a small but meaningful initial manufacturing goal:
Produce 1,000 experimental PhageGuard dressing units as a hypothetical pilot-scale batch.
This would not mean manufacturing 1,000 clinically usable products immediately.
Rather, it would provide a useful engineering scenario for understanding the relationship between biological manufacturing, formulation, and production scale.
📐 3.1 How much would I need?
For a simplified calculation, I would assume:
Target output: 1,000 dressing units.
Nominal dressing area: 5 cm × 5 cm.
Hydrogel equivalent per dressing: 2 mL.
Illustrative phage-containing fraction: 0.5 mL per dressing.
These numbers are engineering assumptions for the homework, not clinically validated specifications.
The total hydrogel volume would be:
1,000 dressings × 2 mL/dressing = 2,000 mL
Total hydrogel equivalent: 2 L
The total phage-containing formulation volume would be:
1,000 dressings × 0.5 mL/dressing = 500 mL
Total phage-containing formulation: 0.5 L
The remaining formulation volume would be contributed by the compatible dressing matrix and other validated components.
The fraction of phage-containing formulation and the biologically active quantity per dressing would ultimately depend on stability, release behavior, and preclinical dose-response studies.
I would not assume that a given phage concentration is therapeutically effective simply because it can be incorporated into the hydrogel.
⏱️ 3.2 How long would manufacturing take?
I would divide the process into three broad stages:
Manufacturing stage
Illustrative planning estimate
Biological production, recovery, and quality testing
5–10 working days
Hydrogel preparation and dressing fabrication
2–3 working days
Packaging, verification, and preliminary release testing
2–5 working days
Total sequential estimate
9–18 working days
These estimates are hypothetical planning assumptions rather than experimentally measured manufacturing rates.
In practice, the timeline would depend on:
Biological yield and batch reproducibility.
Purification and impurity-removal requirements.
Phage stability during processing.
Dressing fabrication throughput.
Quality-control testing and acceptance criteria.
Whether different manufacturing stages can overlap.
For example, if a pilot fabrication system could prepare 100 dressing units per working day, producing 1,000 units would require:
1,000 units ÷ 100 units/day = 10 working days
If the system could prepare 250 units per working day:
1,000 units ÷ 250 units/day = 4 working days
The second scenario illustrates how increasing fabrication capacity could reduce processing time.
However, the throughput numbers are assumed. They would need to be verified experimentally, and increased fabrication speed would not necessarily accelerate biological production or product-release testing.
What changes at larger scale?
An important lesson from HTBAA is that the equipment used to discover a promising formulation may not be appropriate for manufacturing the final product.
At the research stage, small-volume experiments would help evaluate material compatibility, reproducibility, and stability.
At larger scale, the process would require appropriately validated biological production, purification, material manufacturing, and packaging systems.
The challenge would not simply be producing more bacteriophages.
It would be producing consistent, well-characterized, safe, and stable dressing units.
That is the kind of manufacturing problem I would like to understand better throughout the course.
4. 🤖 How would I use automation?
For PhageGuard, I see automation as a way to connect two areas that are often studied separately:
Biological production and biomaterial formulation.
I would not begin by attempting to automate the entire manufacturing process.
Instead, I would use automation to answer smaller, well-defined questions about the compatibility of bacteriophages with different materials and conditions.
🧪 1. Experimental design
The first step would be developing a structured Design of Experiments (DoE) framework.
Rather than choosing one hydrogel formulation and hoping it works, I would compare several material characteristics systematically.
An initial screening matrix could explore:
polymer family × material composition × storage condition × evaluation time
The primary outcomes would include:
Physical integrity of the hydrogel.
Changes in material properties during storage.
Compatibility with a model biological component.
Reproducibility between formulations.
Where authorized phage research is possible, a separate study could investigate whether validated formulations preserve phage activity.
This separation would be useful because it would allow me to learn about the biomaterial even if phage experiments were not available through the course infrastructure.
🤖 2. Automated liquid handling
Desktop liquid-handling robots, such as Opentrons, could help prepare multiple small-volume formulations with greater consistency.
Potential applications include:
Dispensing compatible hydrogel precursors.
Preparing structured experimental matrices.
Creating technical replicates.
Standardizing sample distribution.
Reducing variability in formulation preparation.
For early course-based experiments, I would prioritize noninfectious materials and safe model systems.
Handling bacteriophages and bacterial hosts would require suitable facilities, supervision, and authorization.
📊 3. Automated characterization
Automation could also support repeated measurements of material performance.
Depending on available instruments, useful measurements could include:
💧 Swelling behavior: How much liquid a material absorbs.
🧪 Material stability: Whether the formulation changes over time.
🔬 Biological compatibility: Whether a suitable model biological component maintains its expected properties.
For actual bacteriophage formulations, specialist laboratories would also need to evaluate biological activity and purity using validated methods.
I would not assume that conventional optical absorbance measurements alone can establish phage infectivity or therapeutic potency.
💻 4. From experiments to decisions
I would use Python or Jupyter Notebook to organize and analyze the experimental results.
A simple workflow might look like:
material formulation → measured properties → stability profile → formulation comparison → next experiment
The objective would be to move from subjective observations to quantitative and reproducible design decisions.
For example, a formulation that looks physically attractive but fails to preserve biological activity would not be a successful candidate.
Similarly, a formulation that maintains biological activity but lacks suitable mechanical or storage properties might not become a practical dressing.
I would want to evaluate both sides of the problem.
🔬 5. Could microfluidics help?
HTBAA also introduces programmable digital microfluidics and cell-free manufacturing.
I would be interested in exploring whether small-volume automated platforms could eventually help compare material formulations while reducing reagent consumption.
However, I recognize that not every microfluidic platform is suitable for preparing hydrogels or handling intact phages.
The immediate objective would therefore be to understand which available automation technologies are compatible with the physical and biological requirements of the project.
🌍 Where my node becomes especially important
One reason I am particularly excited about PhageGuard is that my node has previous experience working with bacteriophages.
I think this could make the project especially valuable as a learning opportunity.
I would love to understand:
What types of bacteriophage research have already been conducted at the node?
What biological and experimental challenges did the team encounter?
Which questions would be realistic for a student project?
Could the node help evaluate the feasibility of a phage–biomaterial research approach?
Would it be more useful to begin with phage stability, material compatibility, or biological characterization?
I do not assume that access to the node automatically means I can perform every proposed experiment.
Similarly, HTBAA remote laboratories provide opportunities to learn automation, but they do not necessarily support direct bacteriophage production or infection assays.
I would see the different research environments as complementary:
HTBAA automation → How can I compare formulations systematically?
Node expertise → What can I realistically investigate about bacteriophages?
Biomedical engineering → How can those findings inform the design of a functional material?
My goal would be to develop a project that is scientifically interesting while remaining realistic about available infrastructure, expertise, and biosafety requirements.
5. ❓ What don’t I know yet?
At this stage, PhageGuard is a research concept rather than a validated wound-care technology.
I think its feasibility depends on answering three particularly important questions.
1. 🦠 Can bacteriophages provide sufficiently useful antibacterial coverage?
Bacteriophage specificity is one of the most attractive characteristics of this approach, but also one of its biggest limitations.
A phage may be highly effective against a susceptible bacterial strain while having little or no activity against closely related strains.
This raises several questions:
Which bacterial pathogens should PhageGuard target first?
Would one bacteriophage be sufficient, or would several eventually be necessary?
How could we account for differences between bacterial strains and the possible development of phage resistance?
I would initially focus on one well-defined biological target rather than attempting to develop a universal antimicrobial dressing.
The PhagoBurn clinical trial, which investigated bacteriophage therapy for P. aeruginosa burn wound infections, illustrates both the promise and the difficulties of translating phage biology into a reproducible therapeutic product [4].
I would also need to understand how phage selection, genetic characterization, and safety assessment influence manufacturing decisions.
2. 🧪 Can a biomaterial preserve phage activity during manufacturing and storage?
This may be the most interesting engineering challenge of the project.
A bacteriophage can have excellent antibacterial activity under laboratory conditions but become unsuitable for a final product if it loses activity during incorporation, drying, packaging, or storage.
I would want to investigate:
Which material properties affect biological stability?
Can an appropriately designed matrix preserve phage activity over time?
How do storage conditions influence the formulation?
Can release behavior be controlled without compromising biological function?
Could a practical formulation eventually avoid demanding cold-chain requirements?
This last question is particularly meaningful for rural healthcare.
A material that requires highly controlled storage conditions might be difficult to distribute in remote areas.
However, ambient-temperature stability would need to be demonstrated rather than assumed.
Published work on phage-loaded hydrogels shows that formulation chemistry and storage conditions can substantially influence bacteriophage stability [2,3].
This makes the interaction between the phage and the material a central research problem, not a minor formulation detail.
3. 🩹 Can PhageGuard become a safe, affordable, and clinically meaningful product?
Even if a formulation preserves bacteriophage activity, several important challenges would remain.
The manufacturing process would also need to address possible impurities originating from bacterial production.
Additionally, I would need to understand the regulatory pathway for a product that combines a biologically active agent with a medical dressing.
This is particularly important because PhageGuard would not be equivalent to a conventional passive wound dressing.
The intended antimicrobial action would likely introduce additional therapeutic and regulatory considerations.
From an accessibility perspective, I would also need to investigate the actual cost of:
Producing and purifying bacteriophages.
Preparing the biomaterial.
Performing quality-control testing.
Packaging and distributing the final product.
Maintaining stability during storage and transportation.
A product is not truly accessible simply because its raw materials are inexpensive.
Its complete manufacturing and distribution process must also be feasible.
🌱 The bigger question
Behind all three challenges, there is a broader question that I find especially important:
Can we transform the biological specificity of bacteriophages into a practical biomedical material without losing the accessibility that originally motivated the idea?
PhageGuard would only be worth developing if it could demonstrate a meaningful advantage over existing wound-care technologies for a clearly defined use case.
I also recognize that therapeutic bacteriophages are not a substitute for basic wound hygiene, established infection-prevention practices, appropriate antibiotics, or timely medical assessment.
For that reason, the first realistic milestone would not be a product ready for use by farmers.
It would be a laboratory proof of concept showing that a biocompatible material can preserve the relevant biological properties of a well-characterized bacteriophage, under suitable research conditions.
If that initial question can be answered, future work could investigate antimicrobial performance, manufacturing reproducibility, safety, and clinical relevance.
I want to explore whether bacteriophages, biomaterials, and automated biomanufacturing can come together to create a new approach to accessible wound care.
What makes this project especially exciting to me is the possibility of learning from a node already experienced in bacteriophage research while connecting that knowledge to my background in Biomedical Engineering.
Ultimately, I would love to see biotechnology contribute to practical healthcare solutions for rural communities in Colombia.
That is the motivation I would like to bring into HTBAA.
📚 References
[1] World Health Organization. (2024). WHO bacterial priority pathogens list, 2024: Bacterial pathogens of public health importance to guide research, development and strategies to prevent and control antimicrobial resistance.
[2]Insights into the Preparation of and Evaluation of the Bactericidal Effects of Phage-Based Hydrogels. (2024). Review of phage-containing hydrogel formulations and their potential biomedical applications.
[3]Novel delivery systems for phages and lysins in the topical management of wound infections: A narrative review. (2025). Review of biomaterial-based systems for topical phage delivery.
[4] Jault, P., et al. (2019). Efficacy and tolerability of a cocktail of bacteriophages to treat burn wounds infected by Pseudomonas aeruginosa (PhagoBurn): A randomised, controlled, double-blind phase 1/2 trial. The Lancet Infectious Diseases, 19(1), 35–45.
[5]Current Knowledge and Perspectives of Phage Therapy for Combating Refractory Wound Infections. (2024). Review of bacteriophage therapy and delivery approaches for difficult-to-treat wound infections.